Basis-set convergence
Total energies and most properties improve as you move from small basis sets (minimal, split-valence) toward large ones (triple-zeta with diffuse and polarization). This tutorial shows how to run a family of calculations, tabulate the trend, and decide when you’ve hit the point of diminishing returns.
Before you start
Complete Molecular Hartree-Fock first. Save the blocks below
in one script and run them in order with the installed engine’s Python, from
a directory outside the checkout. The first block defines mol and retains
converged results for reuse by the property and plotting blocks. Coordinates
are in bohr; energies are in hartree. Plotting is optional and needs Matplotlib.
Start with the small basis before attempting the larger points. The sweep
uses the low-level RHF driver, so it does not automatically write the complete
run_job output family. For resource planning, see
direct SCF and memory; for archived studies,
use distinct run_job output stems and retain the matching settings.
A Dunning-family sweep on water
The correlation-consistent series cc-pVXZ (X = D, T, Q)
(Dunning, Thom H. (1989). *Gaussian basis sets for use in correlated molecular calculations. I. The atoms boron through neon and hydrogen.* Journal of Chemical Physics, **90**(2), 1007–1023. doi:[10.1063/1.456153](https://doi.org/10.1063/1.456153)) and its augmented variants
(Kendall, Rick A., Dunning, Thom H., and Harrison, Robert J. (1992). *Electron affinities of the first-row atoms revisited. Systematic basis sets and wave functions.* Journal of Chemical Physics, **96**(9), 6796–6806. doi:[10.1063/1.462569](https://doi.org/10.1063/1.462569)) systematically approach the HF and correlation
complete-basis-set (CBS) limit. Run HF on water at each:
from vibeqc import Atom, BasisSet, Molecule, run_rhf

mol = Molecule([
    Atom(8, [ 0.0,  0.00,  0.00]),
    Atom(1, [ 0.0,  1.43, -0.98]),
    Atom(1, [ 0.0, -1.43, -0.98]),
])

print(f"{'basis':12s}  {'nbasis':>6s}  {'E (Ha)':>16s}  {'ΔE (mHa)':>10s}  {'iters':>5s}")
print("-" * 60)

basis_names = ("sto-3g", "cc-pvdz", "cc-pvtz", "cc-pvqz")
results = {}
previous = None
for name in basis_names:
    basis = BasisSet(mol, name)
    r = run_rhf(mol, basis)
    if not r.converged:
        raise RuntimeError(f"RHF did not converge for {name}; inspect its SCF trace")
    results[name] = (basis, r)
    delta = ""
    if previous is not None:
        delta = f"{(r.energy - previous) * 1000:+10.3f}"
    print(f"{name:12s}  {basis.nbasis:6d}  {r.energy:16.8f}  {delta:>10s}  "
          f"{r.n_iter:5d}")
    previous = r.energy
Illustrative output from the original example (not regenerated here):
basis         nbasis           E (Ha)   ΔE (mHa)  iters
------------------------------------------------------------
sto-3g             7       -74.94956615                  8
cc-pvdz           24       -76.02431388   -1074.748     10
cc-pvtz           58       -76.05605100     -31.737     12
cc-pvqz          115       -76.06394333      -7.892     15
Reading across:
ΔEshrinks by roughly 1/4 with each level up (7.892 / 31.737), classic asymptotic behavior of the correlation-consistent series. Extrapolating to CBS by fitting an exponential in the cardinal number X through the three cc-pVXZ points gives-76.0666 Haat the HF limit.cc-pVTZcaptures 99.3% of the STO-3G to cc-pVQZ improvement for 1/2 the basis functions, a reasonable default for production HF.Timing scales as
O(N^4)in the basis-function count for HF, so doubling the basis multiplies the cost by ~16.
A polarized Pople sweep
A continuation using mol and the imports from the first block:
# doc-audit: skip - continues with mol and imports from the first block
for name in ("6-31g", "6-31g*", "6-31g**", "6-311g**"):
    basis = BasisSet(mol, name)
    r = run_rhf(mol, basis)
    if not r.converged:
        raise RuntimeError(f"RHF did not converge for {name}")
    print(f"{name:14s}  nbasis={basis.nbasis:3d}  E = {r.energy:.6f}")
6-31g         nbasis= 13  E = -75.982973
6-31g*        nbasis= 18  E = -76.006678
6-31g**       nbasis= 24  E = -76.021164
6-311g**      nbasis= 30  E = -76.045115
Adding polarization on heavy atoms (
*) gains ~24 mHa.Adding polarization on hydrogens as well (
**) another ~14 mHa.Going from 6-31 → 6-311 split-valence ~24 mHa.
Diffuse-augmented bases (6-311++g**, aug-cc-pvtz) are
essential for anions, weak-interaction energies, and excited-state
calculations. vibe-qc’s basis library ships them, but their small
exponents can trigger linear-dependency issues in certain tight
geometries. Inspect the overlap diagnostics and follow the
linear-dependence guide. Increasing
max_iter does not repair an ill-conditioned basis.
Properties that converge differently
Total energies are the slowest-converging quantity. Relative energies (reaction energies, barriers) and properties (dipoles, bond orders, frequencies) converge faster because error cancellation helps, but not always monotonically.
The following block reuses the converged Dunning-family results; it does not repeat their SCF calculations. Keep each result paired with its original basis and geometry.
# doc-audit: skip - reuses mol and results from the first block
from vibeqc import mulliken_charges, dipole_moment

for name in ("cc-pvdz", "cc-pvtz", "cc-pvqz"):
    basis, r = results[name]
    q = mulliken_charges(r, basis, mol)
    mu = dipole_moment(r, basis, mol)
    print(f"{name:14s}  E = {r.energy:.5f}   "
          f"q_O(Mull)= {q[0]:+.3f}   μ(D)= {mu.total_debye:.4f}")
Output:
cc-pvdz         E = -76.02431   q_O(Mull)= -0.269   μ(D)= 1.9131
cc-pvtz         E = -76.05605   q_O(Mull)= -0.471   μ(D)= 1.8847
cc-pvqz         E = -76.06394   q_O(Mull)= -0.497   μ(D)= 1.8648
Observe:
Dipole converges cleanly, within 0.03 D by cc-pVTZ, within 0.02 D at cc-pVQZ. The experimental gas-phase value is 1.855 D; cc-pVQZ HF is already on top of it, but this is partially a cancellation of errors (HF over-polarises, no correlation pulls it back) rather than true convergence.
Mulliken charges are a mess, they wander even within a consistent family. Don’t chase a particular value; use them only for within-set comparisons.
Tabulating the trend in a figure
Plot the retained energies against basis-function count without rerunning the sweep. This block continues from the first calculation block:
# doc-audit: skip - reuses basis_names and results from the first block
import matplotlib.pyplot as plt

nbasis = [results[name][0].nbasis for name in basis_names]
energies = [results[name][1].energy for name in basis_names]

fig, ax = plt.subplots()
ax.plot(nbasis, energies, "o-")
ax.set_xlabel("Number of basis functions")
ax.set_ylabel("E(RHF) / Ha")
ax.set_xscale("log")
for n, e, name in zip(nbasis, energies, basis_names):
    ax.annotate(name, (n, e), xytext=(4, 4), textcoords="offset points")
fig.tight_layout()
fig.savefig("h2o_basis_convergence.png", dpi=150)
For a three-point cc-pVXZ extrapolation to the HF CBS limit, fit
E(X) = E_CBS + A * exp(-α X) with all three parameters free (here
α comes out at 1.39); the fixed α ≈ 1.63 of A. Halkier,
T. Helgaker, P. Jørgensen, W. Klopper and J. Olsen, Chem. Phys.
Lett. 302, 437 (1999) serves two-point extrapolations.
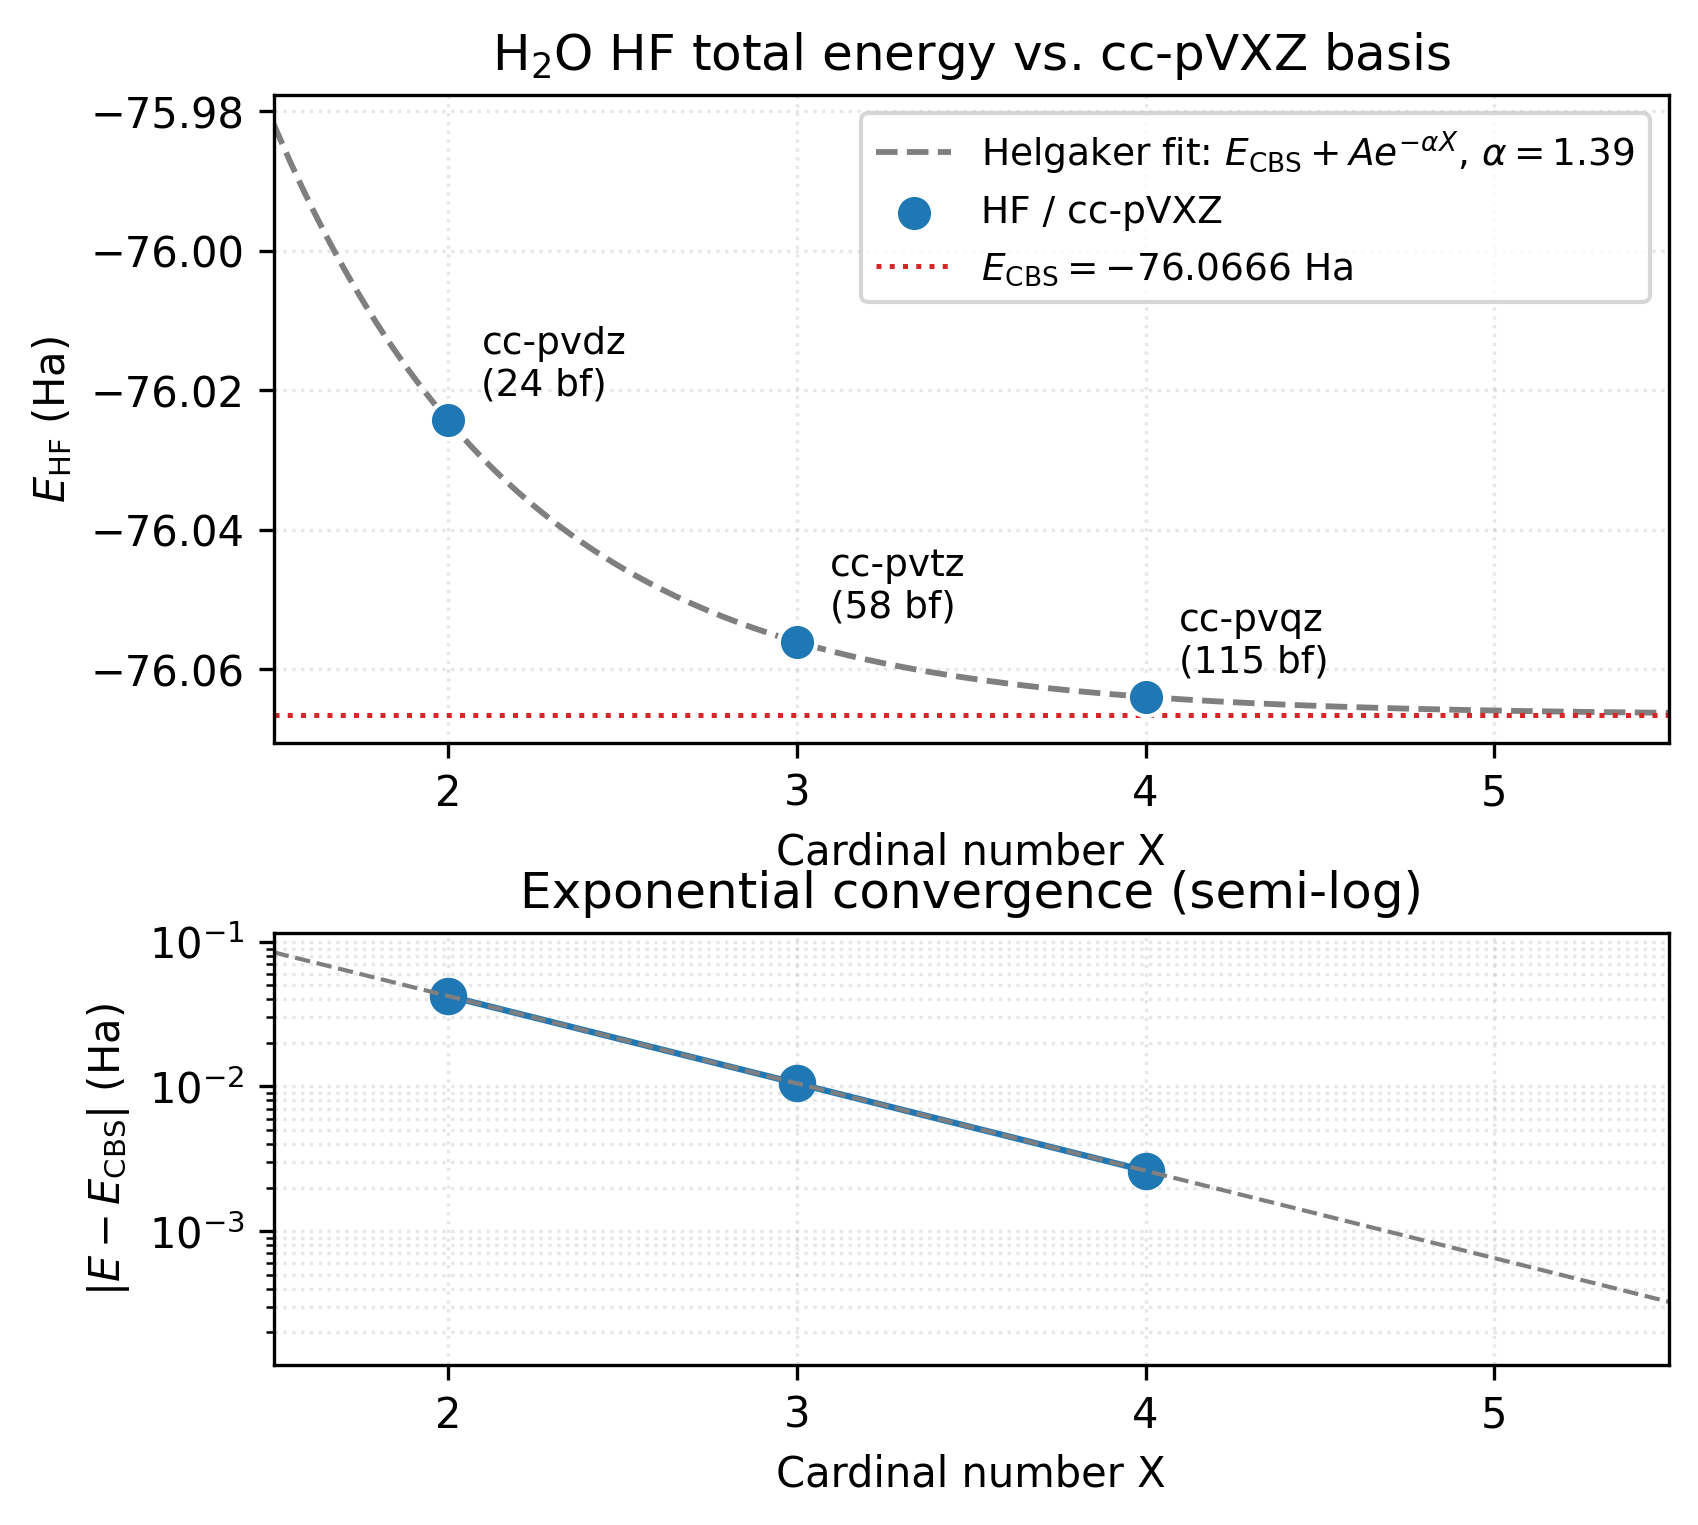
The linked figure illustrates a fit to the recorded cc-pVXZ values. The short plotting block above produces only the energy-versus-basis-count plot; it does not produce the fitted two-panel figure shown here.
A three-point exponential with three free parameters has no remaining data points to test the fit. A smooth curve through those points does not establish the accuracy of the extrapolated limit. Report which cardinal numbers and fit form were used, exclude STO-3G from the cc-pVXZ fit, and distinguish a fitted estimate from a calculated energy. The example’s fitted limit is not an independent numerical reference.
The figure is regenerated by
examples/plots/water-basis-convergence.py.
Notes
Basis-set superposition error (BSSE) matters for interaction-energy calculations on weakly-bound complexes. No counterpoise correction is built in yet, but you can compute it manually by running the monomer calculations in the dimer basis and subtracting.
Basis linear dependencies. At very large bases (
aug-cc-pVQZand bigger, or diffuse functions in close-packed geometries) the overlap matrix can become nearly singular. Inspect removed directions and conditioning warnings; convergence is not guaranteed. Follow the linear-dependence guide. For periodic calculations this is why thepob-*solid-state basis family exists, small-exponent primitives trimmed deliberately.
Theory
The sections below cover the machinery behind the sweeps above: what a basis set is as an expansion in contracted Gaussians, the two common basis hierarchies (Pople and correlation-consistent), Helgaker’s CBS extrapolation formula, what polarization functions buy you, and why solids need their own bespoke basis families.
What a basis set is
In a practical electronic-structure calculation, every molecular orbital is expanded in a fixed, finite set of atom-centered basis functions \(\{\chi_\mu\}\):
With an infinite, complete basis you get the exact answer of whatever method you’re using (HF, DFT, CCSD, …). With a finite basis you get an approximation whose error scales with basis size; the study of that scaling is basis-set convergence.
Each \(\chi_\mu\) in vibe-qc is a contracted Gaussian-type orbital:
a fixed linear combination of \(K_\mu\) primitive Gaussians with pre-determined exponents \(\alpha_{\mu p}\) and contraction coefficients \(d_{\mu p}\), multiplied by a solid harmonic \((\mathbf{r} - \mathbf{R}_A)^{\ell_\mu}\). The contraction saves the SCF driver from having to optimize every primitive separately, the basis-set designer has already done that for atomic ground states and frozen the inner-shell mix.
Two common hierarchies
Pople split-valence (6-31G*, 6-311G**). Inner-shell contracted
to one function, valence split into two (6-31G,
Ditchfield, R., Hehre, W. J., and Pople, J. A. (1971). *Self-Consistent Molecular-Orbital Methods. IX. An Extended Gaussian-Type Basis for Molecular-Orbital Studies of Organic Molecules.* Journal of Chemical Physics, **54**(2), 724–728. doi:[10.1063/1.1674902](https://doi.org/10.1063/1.1674902), and W. J. Hehre, R. Ditchfield and
J. A. Pople, J. Chem. Phys. 56, 2257 (1972), doi:10.1063/1.1677527)
or three (6-311G, Krishnan, R., Binkley, J. S., et al. (1980). *Self-consistent molecular orbital methods. XX. A basis set for correlated wave functions.* Journal of Chemical Physics, **72**(1), 650–654. doi:[10.1063/1.438955](https://doi.org/10.1063/1.438955)).
Polarization functions (Hariharan, P. C. and Pople, J. A. (1973). *The influence of polarization functions on molecular orbital hydrogenation energies.* Theoretica Chimica Acta, **28**(3), 213–222. doi:[10.1007/BF00533485](https://doi.org/10.1007/BF00533485); the
* = d-type on heavy atoms; ** = d and p on hydrogen) give angular
flexibility beyond the valence’s \(\ell\)-range. Diffuse functions (+ on heavy atoms; ++ with H;
Frisch, M. J., Pople, J. A., and Binkley, J. S. (1984). *Self-consistent molecular orbital methods 25. Supplementary functions for Gaussian basis sets.* Journal of Chemical Physics, **80**(7), 3265–3269. doi:[10.1063/1.447079](https://doi.org/10.1063/1.447079)) add
small-exponent primitives for anions / Rydberg states / weak
bonds.
Correlation-consistent (cc-pVXZ). Designed by Dunning (Dunning, Thom H. (1989). *Gaussian basis sets for use in correlated molecular calculations. I. The atoms boron through neon and hydrogen.* Journal of Chemical Physics, **90**(2), 1007–1023. doi:[10.1063/1.456153](https://doi.org/10.1063/1.456153)) to systematically recover correlation energy, with the X = D, T, Q, 5, 6 cardinal number controlling both valence-flexibility and polarization depth together. The key property: the incremental basis-set-error contribution falls off systematically with X (exponentially for the HF energy, as \(X^{-3}\) for the correlation energy), so two successive cardinal numbers extrapolate reliably to the complete-basis-set (CBS) limit.
Helgaker’s CBS extrapolation
The canonical extrapolation of the HF total energy uses the exponential form of Halkier, Helgaker, Jørgensen, Klopper and Olsen (1999, cited above):
with \(\alpha \approx 1.63\) fixed across the Dunning cc-pVXZ series. Correlation energies converge more slowly (electron-electron cusps), and the standard fit is the inverse-cubic power law (T. Helgaker, W. Klopper, H. Koch and J. Noga, J. Chem. Phys. 106, 9639 (1997), doi:10.1063/1.473863; A. Halkier et al., Chem. Phys. Lett. 286, 243 (1998), doi:10.1016/S0009-2614(98)00111-0)
Three consecutive cc-pVXZ points are enough to fit either form.
Polarization functions: what they add
A minimal basis (STO-3G, Hehre, W. J., Stewart, R. F., and Pople, J. A. (1969). *Self-Consistent Molecular-Orbital Methods. I. Use of Gaussian Expansions of Slater-Type Atomic Orbitals.* Journal of Chemical Physics, **51**(6), 2657–2664. doi:[10.1063/1.1672392](https://doi.org/10.1063/1.1672392)) has no
angular-momentum flexibility beyond
what atomic ground states need. Adding a \(d\)-type shell to a heavy
atom, or a \(p\)-type to hydrogen, lets the orbital “polarize” in
response to the bonding environment. Without polarization, oxygen
can’t respond to the electronic asymmetry of the O-H bond and
dipoles under-shoot by 15-25%. The * / ** in Pople names and
the P in pVXZ both denote polarization. For anything past a
qualitative sketch, polarization is not optional.
Why solids use different basis sets
In a crystal, diffuse basis functions on neighboring atoms overlap significantly, because in a crystal, every atom has 6 or 12 near neighbors at bonding distance rather than the 1-4 of a molecule. That pushes the overlap matrix toward singularity and the SCF into linear-dependency territory. Solid-state basis families like pob-TZVP (Peintinger-Vilela Oliveira-Bredow 2013, Peintinger, Michael F., Vilela Oliveira, Daniel, and Bredow, Thomas (2013). *Consistent Gaussian basis sets of triple-zeta valence with polarization quality for solid-state calculations.* Journal of Computational Chemistry, **34**(6), 451–459. doi:[10.1002/jcc.23153](https://doi.org/10.1002/jcc.23153)) are redesigned from molecular bases with the smallest-exponent primitives either removed or replaced by less-diffuse alternatives. See Periodic HF on a 1D H chain for the pob-* motivation.
Resources
~250 MB peak RAM, ~5 s on one core (Apple M2 baseline) for the H₂O scan up to cc-pVQZ; each individual SCF is sub-second. cc-pV5Z and beyond on H₂O hit ~30 s and ~1 GB RAM each, the Fock build is \(\mathcal{O}(N_\text{bf}^4)\) and the basis count grows roughly geometrically in the cardinality.
References
Pople basis sets. W. J. Hehre, R. Ditchfield, and J. A. Pople, “Self-consistent molecular orbital methods. XII. Further extensions of Gaussian-type basis sets for use in molecular orbital studies of organic molecules,” J. Chem. Phys. 56, 2257 (1972). The 6-31G* / 6-31G** papers are in the same series.
Correlation-consistent basis sets, original. T. H. Dunning Jr., “Gaussian basis sets for use in correlated molecular calculations. I. The atoms boron through neon and hydrogen,” J. Chem. Phys. 90, 1007 (1989).
Augmented Dunning basis sets. R. A. Kendall, T. H. Dunning Jr., and R. J. Harrison, “Electron affinities of the first-row atoms revisited. Systematic basis sets and wave functions,” J. Chem. Phys. 96, 6796 (1992).
Helgaker CBS extrapolation. T. Helgaker, W. Klopper, H. Koch, and J. Noga, “Basis-set convergence of correlated calculations on water,” J. Chem. Phys. 106, 9639 (1997).
Exponential HF extrapolation (α ≈ 1.63). A. Halkier, T. Helgaker, P. Jørgensen, W. Klopper, and J. Olsen, “Basis-set convergence of the energy in molecular Hartree-Fock calculations,” Chem. Phys. Lett. 302, 437 (1999).
Inverse-cubic correlation extrapolation. A. Halkier, T. Helgaker, P. Jørgensen, W. Klopper, H. Koch, J. Olsen, and A. K. Wilson, “Basis-set convergence in correlated calculations on Ne, N₂, and H₂O,” Chem. Phys. Lett. 286, 243 (1998).
pob-* solid-state basis sets. M. F. Peintinger, D. V. Oliveira, and T. Bredow, “Consistent Gaussian basis sets of triple-zeta valence with polarization quality for solid-state calculations,” J. Comput. Chem. 34, 451 (2013).
Textbook. T. Helgaker, P. Jørgensen, and J. Olsen, Molecular Electronic-Structure Theory, Wiley (2000). Chapter 8 is the comprehensive treatment of Gaussian basis sets.
Next
A DFT-functional-comparison tutorial using the same sweep pattern is next on the list (scan the XC library instead of basis sets).
The periodic HF tutorial uses
pob-tzvpdirectly because standard basis sets are usually too diffuse for crystals.