Geometry optimization
See also the user-guide reference for the native (ASE-free) optimizer’s full API.
Relax a molecular structure toward a stationary geometry with
run_job(..., optimize=True). The examples below select ASE’s
(Larsen, Ask Hjorth, Mortensen, Jens Jørgen, et al. (2017). *The atomic simulation environment - a Python library for working with atoms.* Journal of Physics: Condensed Matter, **29**(27), 273002. doi:[10.1088/1361-648X/aa680e](https://doi.org/10.1088/1361-648X/aa680e)) BFGS line-search backend explicitly so that the
.traj inspection steps use a known file format. optimizer_backend="auto"
chooses ASE when available and the native backend otherwise; supported
methods and constraints depend on that choice.
Run the scripts in a calculation directory outside the checkout. Native
Atom positions are in bohr, while ASE trajectories use angstrom and eV.
An optimized geometry still needs characterization, usually by vibrational
analysis, before calling it a local minimum; it is not a search over all
conformers.
Water at HF/6-31G*
Starting from an approximate geometry:
from vibeqc import Atom, Molecule, run_job

water = Molecule([
    Atom(8, [0.0,   0.0,   0.0  ]),
    Atom(1, [0.0,   1.60, -1.10]),   # slightly off-equilibrium
    Atom(1, [0.0,  -1.60, -1.10]),
])

result = run_job(
    water,
    basis="6-31g*",
    method="rhf",
    output="water_opt",
    optimize=True,
    optimizer_backend="ase",
    write_molden_file=True,
    fmax=0.05,            # eV/Å; ASE checks the largest per-atom force norm
    max_opt_steps=50,
)
if not result.converged:
    raise RuntimeError("The final electronic calculation did not converge")
The geometry optimizer must reach its own force target before the final
single point. result.converged then describes the electronic result, not
an independent geometry or vibrational-stability verdict. A capped geometry
run raises instead of silently proceeding; see handling a stopped run.
Files produced on a successful run:
water_opt.out, initial geometry, optimizer parameters, final optimized geometry, then the full SCF trace at the minimumwater_opt.molden, MOs at the minimum, for Avogadro / Jmolwater_opt.traj, initial structure and accepted optimization frames (ASE binary); view withase gui water_opt.trajwater_opt.xyz, final coordinates in angstrom;.systemand citation files retain provenance, and.qvfis written by default
Reading the output
After the banner and atom table, water_opt.out contains (vibe-qc
0.17.17 development build):
  Geometry optimization (ASE/BFGS) -> water_opt.traj
  fmax = 0.05 eV/A, max_steps = 50

  Optimized geometry
  Atoms (bohr)
  ------------------------------------------------------------
     1  Z=  8       0.00000000      0.00000000     -0.01145730
     2  Z=  1      -0.00000000      1.42616505     -1.09427135
     3  Z=  1       0.00000000     -1.42616505     -1.09427135
  charge=0  multiplicity=1  n_electrons=10

  iter     energy (Ha)  ...
  (SCF trace at the optimized geometry)
  ...
  converged in 9 iterations; E = -76.0093413306 Ha
The OH bond length and HOH angle at this HF/6-31G* minimum: r(OH) = 0.948 Å, HOH = 105.6° (experimental: 0.957 Å, 104.5°). HF slightly under-shoots the bond and opens the angle.
Parameters
The optimization-specific knobs on run_job control the
force-convergence target, the optimizer step limit, and how the
intermediate SCFs are run:
Argument |
Default |
Meaning |
|---|---|---|
|
0.05 |
Force-convergence tolerance (eV/Å) |
|
200 |
Hard iteration limit for the optimizer |
|
|
|
|
|
Applied to every intermediate SCF; bump |
rhf_options / rks_options forward through to every frame of the
optimization, not just the final single point. This matters for
H-bonded systems and floppy molecules where a trial geometry can be difficult
for SCF. Inspect the electronic trace before changing controls. A larger
max_iter helps only if SCF is progressing; neither it nor damping guarantees
a stable trajectory or the intended electronic state.
A harder case: the water dimer
H-bonded clusters sit on a very flat potential-energy surface, plain
BFGS Hessian extrapolation can take poor trial steps before the quadratic
model is trustworthy. The ASE route uses BFGSLineSearch, but its line search
still depends on consistent energies and forces from converged electronic
calculations. Copy examples/h2o_dimer.xyz into your run directory before
running this separate dimer example:
from vibeqc import InitialGuess, RHFOptions, from_xyz, run_job

dimer = from_xyz("h2o_dimer.xyz", charge=0, multiplicity=1)

rhf_opts = RHFOptions()
rhf_opts.max_iter = 200                   # an iteration budget, not a guarantee
rhf_opts.initial_guess = InitialGuess.SAD

run_job(
    dimer, basis="6-31g*", method="rhf",
    output="water_dimer_opt",
    optimize=True,
    optimizer_backend="ase",
    fmax=0.2,                             # loose target for a flat H-bond surface
    max_opt_steps=25,
    rhf_options=rhf_opts,
)
The deliberately loose fmax=0.2 is for a workflow demonstration. It does
not establish an accurate O-O distance or binding energy, and 25 steps may
be insufficient. Keep the same physical model when testing tighter force
and SCF thresholds, then compare the resulting geometry. For interaction
energies, also assess basis convergence, basis-set superposition error and
electron correlation; see basis convergence.
Visualizing the trajectory
The archived animation below records a water HF/6-31G* example with four BFGS steps (five frames). Its step count is illustrative, not a guarantee for the current input, build or tolerances. The trajectory shows the molecule sliding into its equilibrium geometry while the energy drops by ~156 meV:
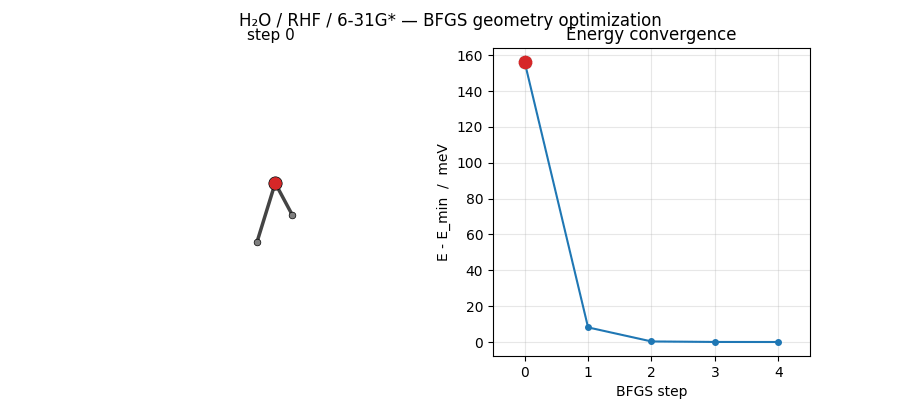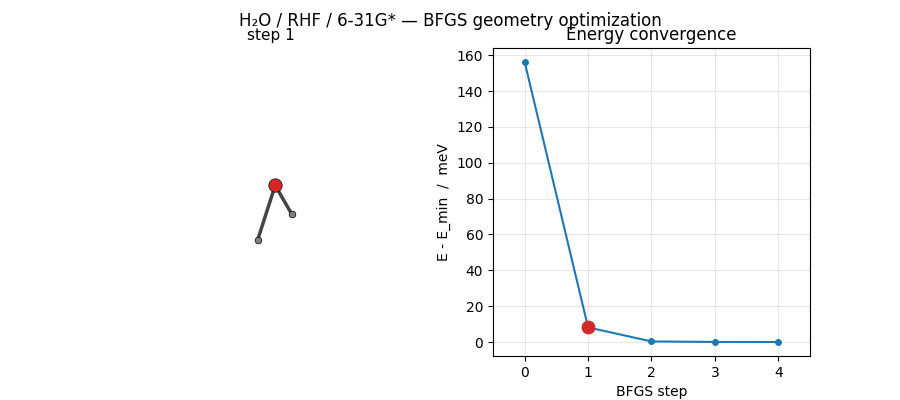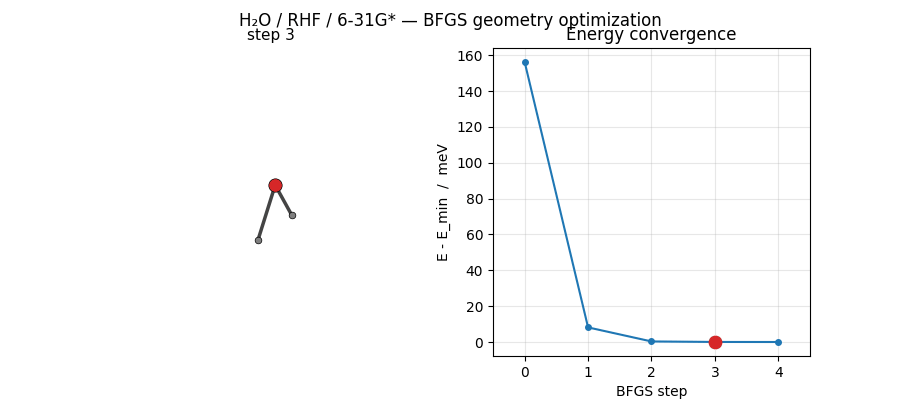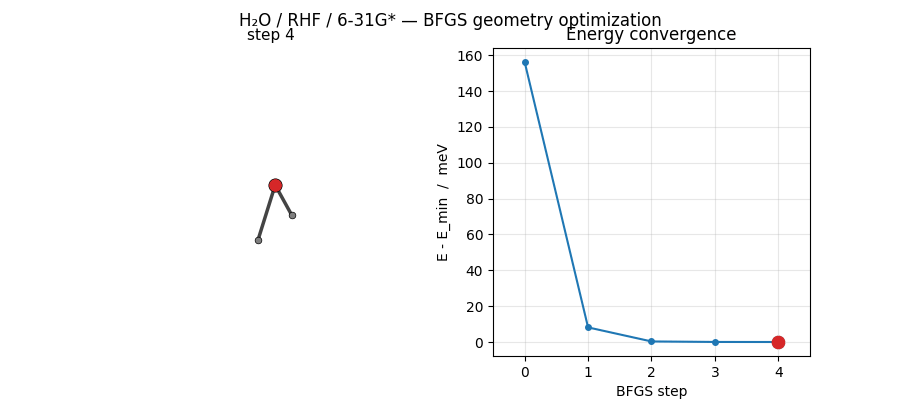
Regenerated by
examples/plots/h2o-opt-trajectory-animation.py
from the per-step .traj file. Pattern is the same as the
Vibrational frequencies:
matplotlib FuncAnimation + PillowWriter, the only difference is
that the input here is a series of saved-during-optimization
positions rather than displaced-along-mode positions.
The .traj file holds positions, energies, and forces for every
frame. ASE’s built-in GUI animates it interactively:
ase gui water_opt.traj
Or inspect programmatically:
import numpy as np
from ase.io import read

frames = read("water_opt.traj", index=":")
if not frames:
    raise RuntimeError("No saved optimization frames")
energies = np.array([f.get_potential_energy() for f in frames])
final_force = np.linalg.norm(frames[-1].get_forces(), axis=1).max()
print(f"Energy dropped {energies[0] - energies[-1]:.4f} eV over "
      f"{len(frames) - 1} saved intervals")
print(f"Final maximum atom force = {final_force:.6f} eV/angstrom")
print(f"Within this example's ASE target: {final_force < 0.05}")
When a run stops before convergence
Read the optimization verdict separately from the SCF verdict. The ASE
route raises a RuntimeError subclass on a step-budget failure; native and
Brent routes raise GeomOptNotConvergedError. Preserve the .out, .xyz
and available trajectory. Restarting from the last geometry starts a new
optimization; it does not restore the previous optimizer’s Hessian history.
Increase the step budget only when the trajectory is progressing toward the same target. Diagnose failed SCFs, inconsistent forces or an unsuitable model before retrying. Loosening the force target changes the acceptance criterion; record that choice instead of treating it as a repair. The user guide describes exception data and low-level return values.
DFT geometries
Reuse water from the first block, not the separate dimer object:
# doc-audit: skip - continues with water from the first example
run_job(
    water, basis="def2-tzvp", method="rks", functional="PBE",
    output="water_pbe_opt",
    optimize=True,
    optimizer_backend="ase",
)
PBE lengthens O-H bonds by ~0.02 Å relative to HF and pulls the HOH angle back toward 104°, generally matching experiment more closely (with the well-known caveat that GGA over-binds H-bonds somewhat).
Brent steepest-descent optimizer
For systems where the Hessian approximation misbehaves (flat potential energy surfaces, dispersion-bound complexes, near-degenerate states), the Brent steepest-descent backend is an alternative based on a one-dimensional line search along the descent direction. It still requires consistent energies and forces and can stop without reaching the requested tolerance. Its line search uses Brent’s method (R. P. Brent, Algorithms for Minimization without Derivatives, Prentice-Hall (1973), chapter 5).
from vibeqc import run_job, Atom, Molecule

mol = Molecule([
    Atom(8, [0.0, 0.00, 0.00]),
    Atom(1, [0.0, 1.43, -0.98]),
    Atom(1, [0.0, -1.43, -0.98]),
])

run_job(
    mol,
    basis="6-31g*",
    method="rks",
    functional="PBE",
    dispersion="d3bj",
    optimize=True,
    optimizer_backend="brent",   # steepest-descent + Brent line search
    output="h2o_pbe_brent",
    output_qvf=True,             # write .qvf archive for vibe-view
    fmax=0.05,
)
Open the animation with vibe-viewer:
vibe-viewer open h2o_pbe_brent.qvf
The QVF archive carries the full trajectory (per-step geometries and energies), the final density, MO metadata, and the system manifest.
The optimizer summary is printed in .out (vibe-qc 0.17.17
development build):
  Geometry optimization (Brent steepest-descent)
  gtol = 9.72e-04 Ha/bohr, max_steps = 200

  Geometry optimization CONVERGED  (Brent steepest-descent, 3 iterations)
  final max|g| = 6.225e-04 Ha/bohr (threshold 9.723e-04)
  the largest gradient component is within the threshold
A run that stops at max_opt_steps before reaching the threshold prints
Geometry optimization NOT CONVERGED instead and ends the job with a
GeomOptNotConvergedError; see
When the optimizer does not converge
in the user guide.
When to use each backend
Situation |
Recommended backend |
|---|---|
Routine organic molecule, tight convergence |
ASE BFGS |
No ASE installed, frozen-atom constraints |
Native L-BFGS-B |
Flat PES (H-bonded clusters, vdW complexes) |
Brent or ASE BFGS |
Near-degenerate states, Hessian misbehaves |
Brent |
Semi-empirical / MLIP methods |
ASE BFGS only (auto-fallback) |
|
Native L-BFGS-B or Brent only: ASE (including the |
Theory
The sections below build up the machinery behind the one-liner: what a stationary point on the potential-energy surface is, where the analytic forces come from, and how the BFGS line-search optimizer uses them to step toward the minimum.
What “optimization” means on a Born-Oppenheimer PES
The Born-Oppenheimer approximation (M. Born and R. Oppenheimer, Ann. Phys. 389, 457 (1927), doi:10.1002/andp.19273892002) decouples nuclear and electronic motion: fix the nuclei at positions \(\mathbf{R} = \{\mathbf{R}_A\}\), solve the electronic Schrödinger equation for those positions, and treat the resulting electronic energy \(E(\mathbf{R})\) as a potential for the nuclei. Geometry optimization is the problem of finding a stationary point on this potential-energy surface (PES):
A local minimum has all positive eigenvalues of the Hessian \(\mathbf{H}_{AB} = \partial^2 E / \partial \mathbf{R}_A \partial \mathbf{R}_B\); a transition state has exactly one negative eigenvalue.
The Hellmann-Feynman theorem and analytic forces
For a variational electronic method at convergence, the forces \(\mathbf{F}_A = -\mathbf{g}_A\) reduce to expectation values of explicit derivatives of the Hamiltonian:
This is the Hellmann-Feynman theorem (Feynman, R. P. (1939). *Forces in Molecules.* Physical Review, **56**(4), 340–343. doi:[10.1103/PhysRev.56.340](https://doi.org/10.1103/PhysRev.56.340)). For an atom-centered Gaussian basis the orbitals themselves move with the nuclei, so the full analytic gradient also needs Pulay forces (Pulay, P. (1969). *Ab initio calculation of force constants and equilibrium geometries in polyatomic molecules. I. Theory.* Molecular Physics, **17**(2), 197–204. doi:[10.1080/00268976900100941](https://doi.org/10.1080/00268976900100941)), corrections from the basis-function derivatives. vibe-qc assembles all the pieces (one-electron derivatives, two-electron ERI derivatives, DFT-grid derivatives for KS, and dispersion-gradient contributions) into an analytic \(\mathbf{g}_A\) at every SCF step. Accuracy is bounded by the SCF tolerance: default settings give forces to a few \(10^{-8}\) Ha/bohr (3 to 4 × \(10^{-8}\) Ha/bohr against a tightly converged SCF for water at HF/6-31G*, vibe-qc 0.17.17 development build).
BFGS: quasi-Newton in quasi-reality
A Newton step along the PES would be
\(\Delta \mathbf{R} = -\mathbf{H}^{-1} \mathbf{g}\), but computing the
Hessian is expensive (the vibrational_frequencies tutorial will do it by finite differences).
Quasi-Newton methods build an approximate inverse Hessian \(\mathbf{B}_k\)
from the history of steps and gradients. BFGS (Broyden-Fletcher-
Goldfarb-Shanno: C. G. Broyden, IMA J. Appl. Math. 6, 76 (1970),
doi:10.1093/imamat/6.1.76; R. Fletcher, Comput. J. 13, 317
(1970), doi:10.1093/comjnl/13.3.317; D. Goldfarb, Math. Comp.
24, 23 (1970), doi:10.1090/S0025-5718-1970-0258249-6;
D. F. Shanno, Math. Comp. 24, 647 (1970),
doi:10.1090/S0025-5718-1970-0274029-X) is the canonical update: given step \(\mathbf{s}_k =
\mathbf{R}_{k+1} - \mathbf{R}_k\) and gradient change
\(\mathbf{y}_k = \mathbf{g}_{k+1} - \mathbf{g}_k\),
Starting from \(\mathbf{B}_0 = \alpha \mathbf{I}\) (a loose diagonal model), BFGS accumulates curvature information so the quadratic model becomes genuinely predictive near the minimum.
The line-search variant used here
vibe-qc’s run_job(..., optimize=True) actually runs ASE’s
BFGSLineSearch, not bare BFGS. The line search enforces the
Wolfe conditions (P. Wolfe, SIAM Rev. 11, 226 (1969),
doi:10.1137/1011036), sufficient-decrease plus curvature, on each
step, so every accepted move reduces the energy. This matters on
flat / weakly-bound surfaces (H-bonded clusters, van-der-Waals
complexes) where plain BFGS’s quadratic model sometimes predicts
wild steps that blow the system apart; the line search catches
them. Cost: 1-3 extra SCFs per BFGS step when the first trial
step is uphill.
Convergence criteria
vibe-qc stops when the maximum-component force is below
fmax (units eV/Å, ASE convention). Default fmax = 0.05 is
“medium-tight” (~1 × 10⁻³ Ha/bohr). Benchmark-quality geometries use
fmax = 0.01; flat H-bonded clusters are often given a looser
fmax = 0.2 for a first pass, because tighter targets cost many
more steps on such a surface.
Resources
~150 MB peak RAM, ~5 s on one core (Apple M2 baseline) for the
H₂O example: 4 BFGSLineSearch steps (5 trajectory frames), each
one HF/6-31G* + analytic gradient. The water-trimer optimization in
examples/molecular/input-h2o-trimer-opt.py is ~5 minutes, basis size
and step count both grow with system size.
References
Born-Oppenheimer approximation. M. Born and J. R. Oppenheimer, “Zur Quantentheorie der Molekeln,” Ann. Phys. 389, 457 (1927).
Hellmann-Feynman theorem. H. Hellmann, Einführung in die Quantenchemie, Deuticke (1937); R. P. Feynman, “Forces in molecules,” Phys. Rev. 56, 340 (1939).
Pulay forces. P. Pulay, “Ab initio calculation of force constants and equilibrium geometries in polyatomic molecules. I. Theory,” Mol. Phys. 17, 197 (1969).
BFGS update. C. G. Broyden, “The convergence of a class of double-rank minimization algorithms,” IMA J. Appl. Math. 6, 76 (1970) and parallel papers by Fletcher, Goldfarb, Shanno (1970). Textbook treatment: J. Nocedal and S. J. Wright, Numerical Optimization, 2nd ed., Springer (2006), chapters 6-8.
Wolfe conditions / line search. P. Wolfe, “Convergence conditions for ascent methods,” SIAM Review 11, 226 (1969).
ASE (the driver). A. H. Larsen et al., “The atomic simulation environment, a Python library for working with atoms,” J. Phys. Condens. Matter 29, 273002 (2017).
Next
Post-SCF analysis, charges, bond orders, and dipole moment at the optimized geometry.
Vibrational frequencies, once the geometry is at a stationary point, compute the Hessian to classify it.