vibe-qc
For renamed products, supported legacy commands, and unchanged package/state identities, see the naming compatibility guide.
Quantum chemistry for molecules and solids.
vibe-qc is a Python + C++17 electronic-structure code. The molecular stack, Hartree-Fock, density-functional theory, Møller-Plesset theory, analytic gradients, D3(BJ) dispersion, is validated against PySCF to machine precision. The periodic stack delivers 1D / 2D / 3D Hartree-Fock and Kohn-Sham DFT with Monkhorst-Pack k-meshes, Ewald-summed Madelung, band structure and density of states, and is growing toward CRYSTAL-style crystalline-orbital calculations at full-SCF accuracy. COOP/COHP bonding analysis and periodic Mayer bond orders are available for 1D/2D/3D systems.
Every calculation leaves behind one file: a QVF archive carrying the structure, the wavefunction, scalar fields, spectra, bands, trajectories, provenance and citations, which vibe-viewer opens in a browser, in a terminal over SSH, as a desktop app, or headlessly from Python. QVF is an open, versioned format with an Apache-2.0 reference writer, meant for the whole quantum-chemistry ecosystem rather than for vibe-qc alone.
Built on libint (Gaussian integrals), libxc (500+ XC functionals), and spglib (crystal symmetry). Licensed MPL-2.0.
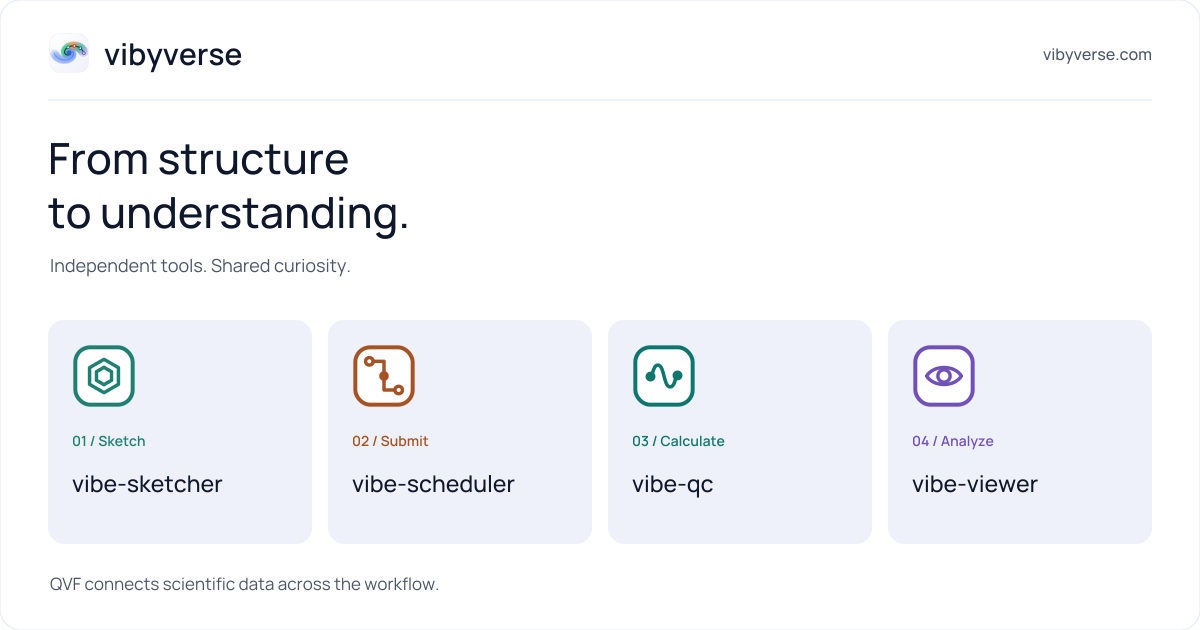
vibe-sketch draws and prepares inputs, vq submits and tracks work, vibe-qc calculates, and vibe-view analyzes results. The workflow is sketch → submit → calculate → analyze; a local calculation can skip queue submission. Install each product independently and follow its own documentation.
Choose where to begin
Your goal |
Start here |
|---|---|
Install the calculation engine |
Start here, then Installation |
Run a first calculation |
|
Open a |
|
Submit, watch, and retrieve queued work |
|
Install or operate a queue host |
|
Choose a method and check its status |
|
Learn methods through worked calculations |
|
Look up options, defaults, and limitations |
|
Browse complete runnable inputs |
|
Diagnose an installation or calculation |
|
Update an existing installation safely |
|
Report a reproducible failure |
|
Develop or document a change |
The Start here page distinguishes the compiled vibe-qc engine from the standalone vibe-viewer package and gives a verification checklist for installing either one or both.
Current source: 0.18.6 - (unnamed)
This manual describes the current source, including changes that may follow the package version shown above. Documentation refreshes do not qualify a new runtime release or widen a method’s validation envelope. See release_process for tagged releases and the branch model.
The repository split makes the viewer, queue, and QVF format independently
adoptable. vibe-basis/ deliberately stays in vibe-qc.
Project |
Version or releases |
Role |
|---|---|---|
vibe-qc |
0.18.6 |
quantum chemistry engine |
vibe-view |
visualization and QVF viewer |
|
vibe-queue ( |
reusable job queue |
|
vibe-basis |
0.14.0 |
basis toolkit, retained here |
QVF |
specification and conformance reference, not a runtime dependency |
See toolset_lifecycle to install or update any of them individually.
💚 vibe-qc is funded by individual sponsors
vibe-qc is built and maintained by one person
in evenings and weekends, on personal hardware, with no institutional
backing. If the project is useful to you - or if you think the Cyclic
Cluster Model reaching CCSD(T) on an open-source code is worth existing -
please consider supporting it via
GitHub Sponsors (recurring
monthly, zero fees) or
Ko-fi (one-time, no GitHub account
required). Every sponsorship directly funds the Claude Max
subscription that drives day-to-day development, the self-hosted
server behind vibe-qc.com, and - most urgently - bigger hardware:
the project currently develops on a single Apple M2 laptop, and that’s
what caps every regression test, CI benchmark, and CRYSTAL-Tutorial
port to small systems. Smallest single actionable item right now:
the NIST Crystal Data SRD 3 single-user
subscription
at $200/year - fully fundable by one sponsor for a year, and
unlocks programmatic access to a curated crystallographic
database for the CCM reference work. See the
support page for the full
pitch and the near-/long-term hardware goals; public sponsors are
listed on the sponsors page.
🖥️ macOS desktop auto-update is not currently supported
The vibe-view desktop app can detect a new release on macOS, but it cannot install the update automatically. Self-update requires Apple Developer registration and Developer ID signing, and that paid registration is currently paywall-gated. Use the manual Download… button for now; see the macOS auto-update sponsorship opportunity for details.
✅ New in v0.18 (Berkelbach’s Badger)
v0.18 is a bounded periodic-correctness release. It freezes the periodic routes and the four MANTA control profiles whose numbers were checked against references, and states the rest as explicit exclusions; the 0.18 scope and the qualified PBC configurations list exactly what is claimed. The v0.18.0 tag was cut on 2026-10-23; its changelog records that full candidate qualification and publication were still pending.
Periodic quadrature. The periodic Becke partition switches atom images off smoothly, so periodic Kohn-Sham energies and weights are continuous in the atomic positions, and the Gamma-point GDF Kohn-Sham gradients now include the motion of that grid.
Periodic ECPs. ECP single points run on the periodic routes, with a source-matched Ewald UKS analytic-gradient preview; other ECP gradients stay refused.
Refusals where a limit does not exist. Dense-cell Gamma RIJCOSX exchange, whose image sum has no limit, is refused; a 1-D wire is modelled as a 3-D cell with vacuum.
Local correlation. DLPNO runs on crystals at Gamma and on BvK-folded meshes, closed-shell DLPNO-CCSD gains LMP2 pair natural orbitals and a compiled pair-screening kernel, and open-shell pair screening defaults to 0.25 tcut_pairs, which changes open-shell energies.
Also in the 0.17.18 to 0.18.0 line: QVF toolkit 0.5 interoperability with canonical excited-state, NTO and PDOS sections; corrected TDDFT transition-dipole factors; native D4 optimisation and gradients; COSMO-RS infinite-dilution activity coefficients; K-commensurate hexagonal k meshes; sparse MRCI; Lebedev rules up to order 95; standalone macOS and Linux distribution builds.
⚠️ MANTA, AICCM/CCM, chi, SECCM and native periodic correlation stay experimental. MANTA’s component certificates bound what they name, not a relaxed total energy.
The preceding v0.17 line (Tew’s Tern) laid the symmetry and periodic
groundwork and the shared operator contract; v0.16 (Pople’s Puffin)
carried the repository split into vibe-qc, vibe-view, vibe-queue and
qvf.
🔜 Current focus after 0.18
The broader production goal for periodic HF, DFT and hybrids, including CRYSTAL23 accuracy and time-to-solution comparisons, is the next theme; the CRYSTAL23 capability matrix shows where each route stands. MANTA continues alongside libint, and native periodic local correlation is deferred to a later feature release. The vibe-qc research paper is under peer review: read the ChemRxiv preprint.
See the roadmap for the full plan.
For the current open-issues list (workarounds, regression-test
pointers, status of each known bug), see
troubleshooting and the
issue tracker.
Periodic COOP/COHP is computed with the same spin Hamiltonian that produced the analysed bands on the GDF routes, and the kernel refuses any other operator; on BIPOLE, AICCM and external-XC routes the DOS and COOP/COHP sections are skipped with a stated reason. Property output on ECP results is tracked as issue 32 on the issue tracker (monorepo number 741); Hirshfeld charges on ECP results now build their promolecule from valence-only free atoms. GDF spectral output retains all accepted eigenstates and uses the requested Gaussian width. If distant states would exceed the property memory limit on one uniform energy grid, it uses nonuniform energy coordinates with resolved Gaussian neighborhoods and exact zeros across empty gaps. The 128 MiB default limit still applies; a genuine refusal is recorded in the run manifest.
Install
Clone the public source over HTTPS; no account or deploy key is required:
git clone https://github.com/vibyverse/vibe-qc.git
Authorized maintainers use their separately configured upstream remote. Private access recipes belong in the external operations documentation.
Then enter the new checkout and install:
cd vibe-qc
./scripts/install.sh --dev                 # native deps + venv + pip install + banner
install.sh accepts --dev (main), --branch NAME (any branch
or tag), and other knobs, see installation.md
for the full surface and the manual setup_native_deps.sh recipe.
git clone selects the project’s default branch, main. The command above
keeps the development line with --dev; the flag-free installer selects
release when that branch exists on the selected origin, or its newest
stable vX.Y.Z tag otherwise. Use --branch vX.Y.Z to pin an exact
published tag for reproducibility.
Public source repositories are listed in repositories and downloads, including each project’s tagged source downloads.
setup_native_deps.sh builds and installs every native dependency
(libint, libxc, spglib, FFTW3, libecpint) into third_party/ and
populates the bundled basis library. Re-running is a no-op if
everything’s already built. Full per-platform dependency lists
(macOS / Arch / Manjaro / Debian / Ubuntu) are in
installation.
Your first calculation
Make a working directory outside the repo, vibe-qc writes its
outputs into the current working directory, and you don’t want
.out / .molden / .traj files landing inside the source
tree:
mkdir -p ~/vibeqc-runs/water
cd ~/vibeqc-runs/water
Save the following as water.py in that directory (any filename
works, vibe-qc just runs whatever Python you point it at):
from vibeqc import Atom, Molecule, run_job

mol = Molecule([
    Atom(8, [ 0.0,  0.00,  0.00]),
    Atom(1, [ 0.0,  1.43, -0.98]),
    Atom(1, [ 0.0, -1.43, -0.98]),
])

run_job(
    mol,
    basis="6-31g*",
    method="rks",
    functional="PBE",
    dispersion="d3bj",
    optimize=True,
    output="water",
)
Run it with the virtual-env’s Python (the one pip install
populated above, not your system python3). Since you’re no
longer in the repo, give the full path:
~/path/to/vibe-qc/.venv/bin/python water.py
Replace ~/path/to/vibe-qc/ with wherever git clone landed.
That ~3-second run writes a set of files into ~/vibeqc-runs/water/.
The four you will reach for first:
water.out, banner, SCF trace, energy breakdown, orbital table, HOMO-LUMO gap, Mulliken / Löwdin charges, Mayer bond orders, dipole, and wall-clock timings.water.qvf, the whole calculation as one archive: structure, optimization trajectory, wavefunction, atom properties, bond orders, SCF history, the run record and the citations. Open it with vibe-viewer, in a browser, in a terminal, or from Python. This one is written by default; passoutput_qvf=Falseto skip it.water.molden, molecular orbitals for Avogadro / Jmol.water.traj, ASE trajectory for the optimization, viewable withase gui water.traj.
The run also leaves .xyz, .bibtex / .references (the
citations for the level of theory you used), .population.txt /
.population.json, and a .system manifest. See
output files for the full family.
Tip
Skip the path prefix. Activate the venv once per shell session and the path resolves automatically - works from any directory:
source ~/path/to/vibe-qc/.venv/bin/activate    # bash / zsh
python water.py                                # uses the venv's python
Deactivate with deactivate when you’re done.
Common mistake: ModuleNotFoundError: No module named 'vibeqc'
means you ran the wrong Python. Either give the full
~/path/to/vibe-qc/.venv/bin/python path or activate the venv
first. The bare .venv/bin/python shorthand only works when your
shell is sitting inside the repo.
See quickstart for a 30-minute end-to-end
walkthrough (HF, periodic SCF, orbital cube), running
for the full “how to invoke vibe-qc scripts” reference (venv,
threading, output capture, SSH workflows),
good practices for the working conventions
nobody tells you (file layout, naming, when to trust a number),
the tour for the wider API surface (run_job, ASE
Calculator, logging, custom basis sets, tests), or dive into the
tutorials for worked examples.
See your results: QVF and vibe-viewer
The water.qvf your first job just wrote is the whole calculation in one
typed, checksummed archive. vibe-viewer is the viewer
for it, and it does not need vibe-qc: it is a wheel-based Python package, so
installing its browser, terminal, and headless modes takes one checkout
installer in the separate viewer repository, with no compiler or native build.
The distribution is not on PyPI yet. The Electron desktop shell is a separate
source-checkout component today.
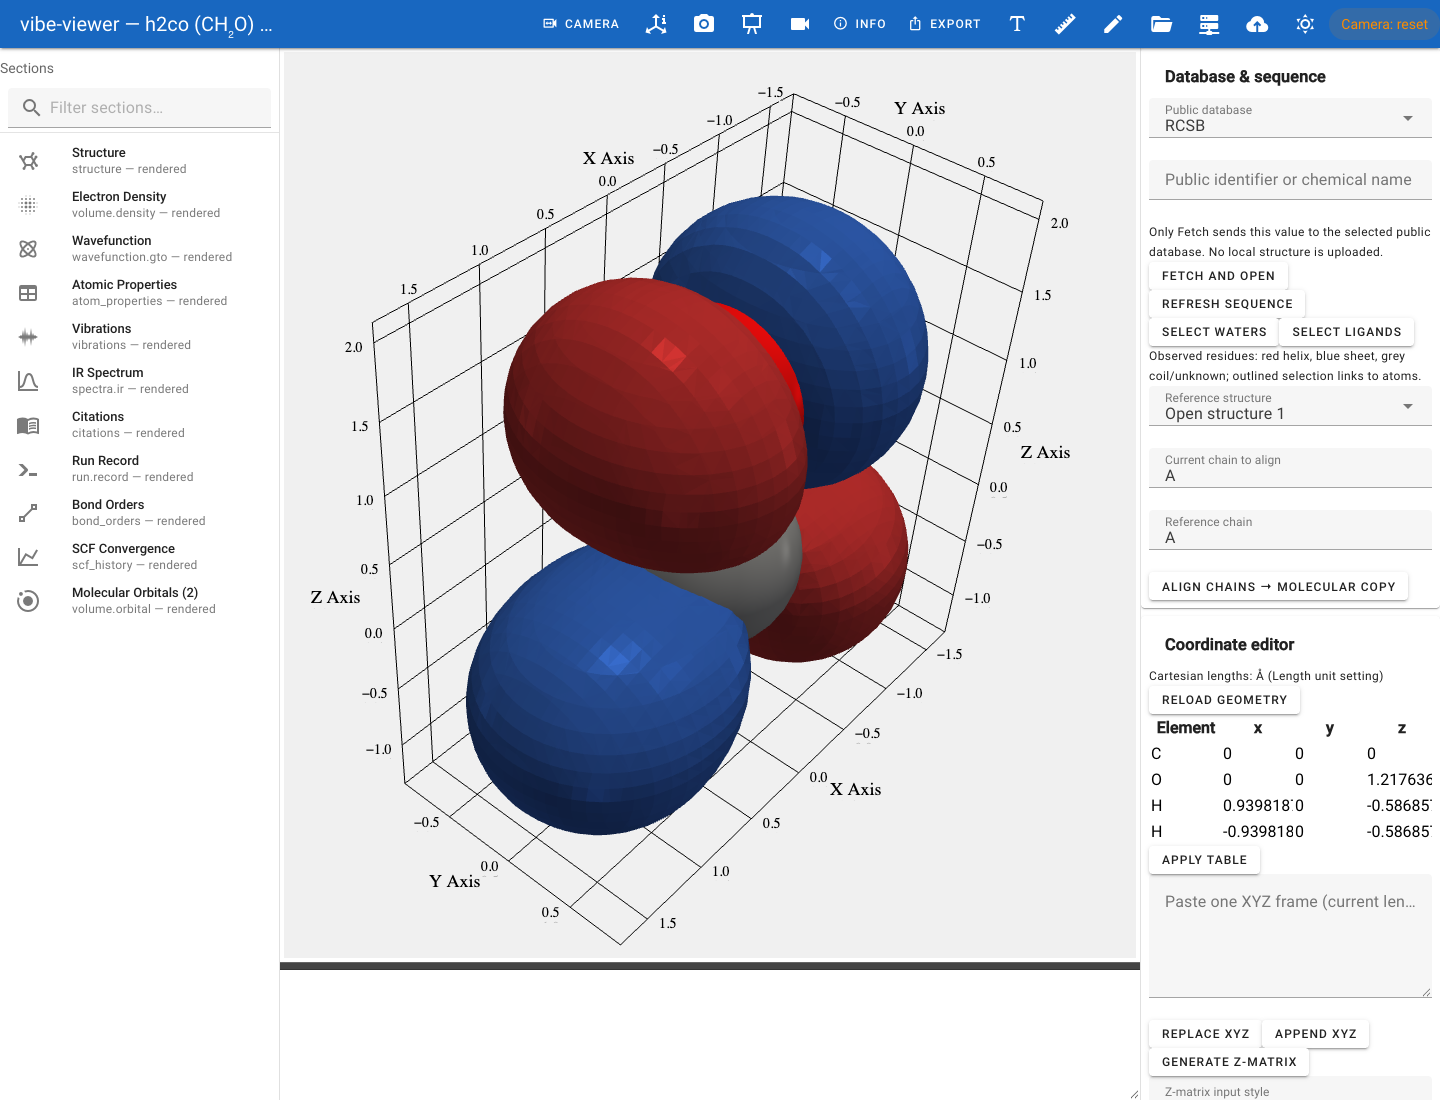
vibe-viewer in the browser: the section browser on the left, the GPU-accelerated 3D viewport in the middle, per-section controls on the right. Here a molecular orbital with both signed lobes.
Four ways to read the same archive, so the environment never dictates whether you can look at your results:
Command |
Needs |
|
|---|---|---|
Browser |
|
an OpenGL context |
Terminal |
|
terminal profile, no display server, no X forwarding |
Desktop app |
|
a source checkout and Electron |
Headless / Python |
|
nothing beyond the base install |
New here? Getting started with vibe-viewer alone installs just the viewer on Linux or macOS. The current checkout creates a built-in demo; the hosted wheel opens files you already have. One wheel carries the released Python viewer modes, with no compiler and no vibe-qc; desktop mode uses the source checkout: Companion release artifacts.
Want the tour? vibe-viewer: an end-to-end walkthrough goes panel by panel.
On a compute node? Terminal mode renders structures, isosurfaces and charts as Unicode braille over SSH.
Writing your own code? QVF is a published, versioned format with a normative specification, a JSON schema, reference writers in Python and zero-dependency C++17, a validator and a conformance corpus, distributed under Apache 2.0 so it can be vendored into any code including a proprietary one. See The QVF format toolkit and Adopting QVF in your own code.
Everything about the format and the viewer lives in one place: QVF and vibe-viewer.
Capabilities today
Molecular. Restricted and unrestricted HF / KS-DFT with analytic nuclear gradients. The full libxc functional library, LDA, GGAs, hybrids, the τ-dependent meta-GGA family (TPSS, M06-2X, SCAN, r²SCAN, r²SCAN01), range-separated hybrids (ωB97X, ωB97X-D, and the VV10-paired ωB97X-V / ωB97M-V), and the PW1PW weighted-sum functional, all supported. Møller-Plesset theory (MP2 / UMP2 / RI-MP2 / SCS-MP2 / SOS-MP2 / open-shell UMP2) and the B2PLYP / DSD-PBEP86 / revDSD-PBEP86-D4 double hybrids. The dense correlated routes are accuracy references for the demonstrated small-molecule envelope; they are not yet claimed as production-size solvers. Density fitting with the RIJK and RIJCOSX Fock-build kernels (RIJCOSX energies within 0.26 mHa of ORCA 6.1.1 on glycine/def2-TZVP). Effective core potentials for heavy-element chemistry. CPCM / COSMO implicit solvation via the SolutePotentialProvider seam. Grimme D3(BJ) / D4 dispersion. 255 bundled Gaussian basis files, including the solid-state pob-* family. Numerical anchors use out-of-process PySCF and ORCA comparisons where documented.
Wavefunction methods. Canonical CCSD(T), the gold-standard
molecular correlation reference: closed-shell, plus open-shell
(UCCSD / UCCSD(T) on a UHF reference, ROHF-reference CCSD / CCSD(T)) and
frozen natural orbitals for cheaper (T). DLPNO-CCSD(T), the
near-linear-scaling local CCSD(T) route, with retained sub-kcal evidence for
explicitly pinned historical recipes (not yet a claim for the current
chemical-core/NormalPNO default), plus open-shell DLPNO-UMP2 and
DLPNO-UCCSD(T) (full reduced-scaling local solver, pilot-anchored against
the dense spin-orbital oracle). DLPNO-(T1) restores off-diagonal localised-Fock
coupling and reaches canonical (T) in the full-domain limit. Multi-root CASCI with state-averaged
CASSCF (analytic nuclear gradient FD-tight
to ~1e-7, shipped in v0.15.0). TDDFT via the Casida linear-response
formalism and the Tamm-Dancoff approximation (RHF/RKS/UHF/UKS), with
Natural Transition Orbitals and FD excited-state gradients.
Green’s-function quasiparticle ionisation energies (GF2 / OVGF,
renormalised GF2). An eigensolver framework (Davidson, LOBPCG, plus
experimental Jacobi-Davidson and GPLHR) selectable by keyword. The
vibeqc.solvers family, Full CI, selected CI, variational 2-RDM,
transcorrelated CI and an experimental DMRG toy solver, via
vibeqc.solvers. General atomisation
energies and RRHO thermochemistry.
Semiempirical + MLIP. The MSINDO semiempirical engine covers
elements H-Xe (Z 1-54), supports NDDO mode for the main group, molecular
geometry optimisation, implicit solvation (COSMO), NEB, velocity-Verlet
molecular dynamics, well-tempered metadynamics, and penalty-function MECI
conical-intersection optimisation; it is production within that scope.
DFTB0 and SCC-DFTB ship as screening and preoptimisation routes (preview:
in-house parameters, no DFTB+ parity). GFN2-xTB and PM6 are experimental,
OM1 to OM3 are experimental and scheduled for retirement, and PM7 is gated.
The semiempirical cyclic cluster model (SECCM) is experimental. The MACE
machine-learning interatomic potential (method="mace") is an external
pre-trained model available as an alternative energy surface. Every label
is defined in Methods: theory and status.
Periodic. 1D / 2D / 3D PeriodicSystem geometry, Monkhorst-Pack
k-meshes with IBZ reduction. Native Gaussian density fitting (GDF,
Γ-point RHF / RKS + hybrids, multi-k KRHF / KRKS; µHa parity vs PySCF
on LiH) and the production GPW (Gaussian Plane Waves) route through
run_periodic_job (Γ-only RHF / UHF / RKS / UKS, multi-k pure-DFT
RKS / UKS; ROHF / ROKS are maintained previews and the MPI grid overlay is experimental). The 3D BIPOLE route covers
Γ + multi-k RHF / UHF / RKS / UKS with the exact Ewald-J split.
Finite-difference forces are the production path; analytic gradients are
a gated preview. The quartet multipole prototype is fail-closed and is not
part of production energies or gradients. 3D Ewald uses a shared α gauge.
Fermi-Dirac, Methfessel-Paxton, and Marzari-Vanderbilt smearing for
metals, with Anderson / Broyden / Kerker density mixers and an AUTO
k-point and smearing recommender. MSINDO periodic CCM through bulk
MgO (Wigner-Seitz, periodic INDO, Ewald Madelung). Experimental
ab-initio CCM: two independent lines. The Γ-CCM line
hosts the neutral finite-torus construction (variants real-gamma and
neutral-bloch) and the 2014 union-and-weight four-centre lineage
(four-center), HF→CCSD(T); χ-CCM is the
finite-character construction (3D SCF + finite-torus correlation). They are
under active side-by-side study on a 28-system benchmark, but no
cross-approach delta is currently reportable; every driver warns. Band structure
and density-of-states plotters. COOP/COHP bonding analysis
(Crystal Orbital Overlap/Hamilton Population; C++ kernels, QVF,
plotters, vibeqc coop CLI). Periodic Mayer bond orders
(k-space generalisation). Fat bands (Mulliken-projected band
weights). QTAIM topological analysis (critical-point search +
bond-path tracing). Gaussian cube + extended-XYZ +
POSCAR + XSF / BXSF writers, plus POSCAR and Extended-XYZ readers
and CIF readers with opt-in geometry symmetrisation.
Tooling. OpenMP parallelism throughout; optional mpi4py
substrate for future MPI parallelism (GPW grid overlay experimental;
production strategy in MPI Parallelization).
Pre-flight memory budget estimator.
ASE Calculator integration for geometry optimization, vibrational
frequencies, and NEB. Automatic per-job citation files
(.bibtex / .references). The
vibe-view interactive 3D viewer for
structure / orbitals / densities / bands / COOP/COHP / spectra /
trajectories from supported QVF sections. The current viewer also displays
reaction schemes and preview images from vibe-sketch. Current viewer source
also renders excited_states tables in the browser, terminal and CLI, with
energies, strengths and orbital contributions. Excited-state energies label
NTOs; the
viewer capability guide
describes support by surface. The
basis_toolkit for
basis-set import/export (BSE, CRYSTAL, G94, NWChem, ORCA). The
vq queue for remote job submission.
See Methods: theory and status for what each method computes and which of the four status labels (production, preview, experimental, gated) it carries, the feature matrix for the option tables, and the roadmap for what’s next.
Where to go next
Getting started
- Start here
- Installation
- Choose the installation you need
- Request repository access
- Clone the source
- Repositories and downloads
- What a successful install does
- Choose the component you need
- Requirements
- One-shot bootstrap
- Per-platform: install the system tools
- What each step does
- Verifying
- Optional Microsoft SKALA-1.1 functional
- Optional terminal viewer (MolTUI)
- Optional basis-set optimization driver (vibe-basis)
- Common issues
- Updating
- Picking a build to test against
- Optional OpenTrustRegion orbital optimizer
- Distribution builds
- Install and maintain the vibe toolset
- Quickstart, your first 30 minutes
- Using Claude to drive vibe-qc
- Setup: point Claude at the docs (the key step)
- A first session, end-to-end: corundum (α-Al₂O₃)
- A second session, Claude Code in the loop: water dimer with dispersion
- Workflow patterns
- What to paste back to Claude
- What Claude can’t see
- Pitfalls and how to dodge them
- End-to-end worked example: SMILES to IR spectrum
- Best practices
- Where to next
- Running vibe-qc
- Choose how to run
- Use the correct Python
- Keep calculations outside the source checkout
- Understand a completed run
- Monitor a foreground run
- Control threads and memory
- Run interactively
- Leave a workstation job running
- Submit through vq
- Submit to Slurm or another HPC scheduler
- Verify an installation or a development change
- Quick diagnosis
- Where to go next
- QCSchema molecular interchange
- Good practices
- Troubleshooting
- Triage the failure first
- Check the environment before rebuilding
- A route refused the request
- SCF didn’t converge
- “Canonical orth dropped too many basis directions”
- Memory abort before the SCF starts
KeyError: 'Ne'(or any other element) on basis load- “Explicit block-mode GAPW is Ha-scale wrong on a molecule like H₂O”
- “DF gradient disagrees by ~115 mHa/bohr”
- “Analytic RHF gradient is wrong”
- “Periodic (BIPOLE) forces / geometry optimisation look wrong”
- CASPT2/NEVPT2
compute_corr_grad=Truegradient has no PT2 correction (v0.15.17) - Multi-k GDF HF / hybrid-KS exchange wrong on diffuse bases in tight cells (v0.15.26 to v0.15.31)
- Multi-k BIPOLE RHF raises
TypeErrorinstantly; BIPOLE RKS ignoresbz_integration="gilat"(v0.15.23 to v0.15.30) ImportError: cannot import name 'EEQOptions' from 'vibeqc._vibeqc_core'ModuleNotFoundError: No module named 'vibeqc'- “Cell-list construction returned 0 cells”
- DFT / COSX wrong on a basis imported as Cartesian via
basis_toolkit(v0.15.0 to v0.15.114) vibeqc-cite: manifest missingafter a job- Native D4 dispersion (
backend="native") scope + accuracy - Things that are not errors
- BIPOLE absolute energies on tight ionic crystals (fixed, corrected gauge is the default at Γ and multi-k, all four drivers)
- Gamma GDF refuses a legacy fallback for a tight-core cell
- Bulk GDF refuses the one-electron memory reservation
- Range-separated GDF reciprocal admission refuses a calculation
- A one-dimensional cell with a k mesh fails on GDF (“no consistent Coulomb gauge for a dim = 1 cell”)
- Prepare a reproducible report
- Updating vibe-qc
- Moving from the archived monorepo
- The easy button
- Variants
- Build-pressure safety
- Advanced diagnostic outline
- Switching between releases
- Going to bleeding-edge
main - Common issues
- Updating an existing development or release installation
- Long-running calculations across an update
- Read-only health check
- Where to go next
- Portable cluster builds
- Tour of the vibe-qc API
- Choose an entry point
- 0. The simple way:
run_job - 1. Load a molecule
- 2. Hartree-Fock energy
- 3. Density Functional Theory (DFT)
- 4. ASE Calculator, the easy path
- 5. Logging the SCF
- 6. Geometry optimization + visualization
- 7. Custom basis sets
- 8. Accessing the full API
- 9. Check a change with the relevant tests
QVF and vibe-view
Tutorial
Design documents -- architecture overviews
- Design - QVF container format for visualization data
- Design,
vibeqc.output(unified output, logging, and citation surface) - Shared symmetry infrastructure
- Symmetry capabilities and production-reduction status
- Integrals first: the public integral primitives and their backends
- MPI Parallelization Strategy
- vibe-qc GPW/GAPW vs GPAW Program – Architectural Comparison
Design documents -- subsystem implementation notes
- QVF-Basis – Architecture Review & Design Decision
- Design, native vibe-qc GDF (Method 1 of the periodic JK roadmap)
- Design, RSGDF 3-centre long-range half (
rsgdf_lr_3c_tensor) - DLPNO-CCSD(T) as one engine: the correlation-source contract for molecules and crystals
- Design,
basis.aosection for basis-function visualization - Design, periodic GAPW / GPW (and a path to PAW + ACE)
- Design: Mixed Density Fitting (MDF) for all-electron µHa periodic GDF
- Design, periodic smearing as a shared backend-agnostic utility
- Design, COSMO-RS thermodynamics from
run_job - χ-CCM / aiccm2026dev-b: an independent finite-torus derivation
- Shared symmetry architecture and production-reduction gates
- Literature-to-code assessment, October 2026
- Γ-CCM (
aiccm2026dev-a) dense four-center scalability - analysis & strategy - Mixed-boundary (wire/slab) Green’s function for the low-D CCM four-center
- Analytic WSSC gradient (Γ-CCM line, four-centre forces): design and implementation spec
- QVF-Basis – Developer README
- QVF-Basis – Conversion Fidelity Matrix
Reference
Project
- Unreleased
- [v0.18.6] - 2026-10-30
- [v0.18.5] - 2026-10-29
- [v0.18.4] - 2026-10-28
- [v0.18.3] - 2026-10-27
- [v0.18.2] - 2026-10-26
- [v0.18.1] - 2026-10-25 - Berkelbach’s Badger
- [v0.18.0] - 2026-10-23 - Berkelbach’s Badger
- [v0.17.26] - 2026-10-22 - Tew’s Tern
- [v0.17.25] - 2026-10-21 - Tew’s Tern
- [v0.17.24] - 2026-10-20 - Tew’s Tern
- [v0.17.23] - 2026-10-19 - Tew’s Tern
- [v0.17.22] - 2026-10-18 - Tew’s Tern
- [v0.17.21] - 2026-10-17 - Tew’s Tern
- [v0.17.20] - 2026-10-16 - Tew’s Tern
- [v0.17.19] - 2026-10-15 - Tew’s Tern
- [v0.17.18] - 2026-10-14 - Tew’s Tern
- v0.17.17 - 2026-10-13 - Tew’s Tern
- [v0.17.16] - 2026-10-12 - Tew’s Tern
- [v0.17.15] - 2026-10-01 - Tew’s Tern
- [v0.17.14] - 2026-10-01 - Tew’s Tern
- [v0.17.13] - 2026-10-01 - Tew’s Tern
- [v0.17.12] - 2026-09-30 - Tew’s Tern
- [v0.17.11] - 2026-09-30 - Tew’s Tern
- [v0.17.10] - 2026-09-26 - Tew’s Tern
- [v0.17.9] - 2026-09-25 - Tew’s Tern
- [v0.17.8] - 2026-09-23 - Tew’s Tern
- [v0.17.7] - 2026-09-22 - Tew’s Tern
- [v0.17.6] - 2026-09-17 - Tew’s Tern
- [v0.17.5] - 2026-09-16 - Tew’s Tern
- [v0.17.4] - 2026-09-16 - Tew’s Tern
- [v0.17.3] - 2026-09-13 - Tew’s Tern
- [v0.17.2] - 2026-09-13 - Tew’s Tern
- [v0.17.1] - 2026-09-10 - Tew’s Tern
- [v0.17.0] - 2026-09-09 - Tew’s Tern
- [v0.16.1] - 2026-09-08 - Pople’s Puffin
- [v0.16.0] - 2026-09-08 - Pople’s Puffin
- [v0.15.162] - 2026-09-06 - Neese’s Cheetah
- [v0.15.161] - 2026-09-05 - Neese’s Cheetah
- [v0.15.160] - 2026-09-04 - Neese’s Cheetah
- [v0.15.159] - 2026-09-03 - Neese’s Cheetah
- [v0.15.158] - 2026-09-02 - Neese’s Cheetah
- [v0.15.157] - 2026-09-02 - Neese’s Cheetah
- [v0.15.156] - 2026-08-31 - Neese’s Cheetah
- [v0.15.155] - 2026-08-30 - Neese’s Cheetah
- [v0.15.154] - 2026-08-29 - Neese’s Cheetah
- [v0.15.153] - 2026-08-29 - Neese’s Cheetah
- [v0.15.152] - 2026-08-28 - Neese’s Cheetah
- [v0.15.151] - 2026-08-28 - Neese’s Cheetah
- [v0.15.150] - 2026-08-28 - Neese’s Cheetah
- [v0.15.149] - 2026-08-28 - Neese’s Cheetah
- [v0.15.148] - 2026-08-27 - Neese’s Cheetah
- [v0.15.147] - 2026-08-27 - Neese’s Cheetah
- [v0.15.146] - 2026-08-27 - Neese’s Cheetah
- [v0.15.145] - 2026-08-27 - Neese’s Cheetah
- [v0.15.144] - 2026-08-26 - Neese’s Cheetah
- [v0.15.143] - 2026-08-26 - Neese’s Cheetah
- [v0.15.142] - 2026-08-25 - Neese’s Cheetah
- [v0.15.141] - 2026-08-25 - Neese’s Cheetah
- [v0.15.140] - 2026-08-25 - Neese’s Cheetah
- [v0.15.139] - 2026-08-24 - Neese’s Cheetah
- [v0.15.138] - 2026-08-21 - Neese’s Cheetah
- [v0.15.137] - 2026-08-19 - Neese’s Cheetah
- [v0.15.136] - 2026-08-18 - Neese’s Cheetah
- [v0.15.135] - 2026-08-17 - Neese’s Cheetah
- [v0.15.134] - 2026-08-14 - Neese’s Cheetah
- [v0.15.133] - 2026-08-14 - Neese’s Cheetah
- [v0.15.132] - 2026-08-14 - Neese’s Cheetah
- [v0.15.131] - 2026-08-12 - Neese’s Cheetah
- [v0.15.130] - 2026-08-12 - Neese’s Cheetah
- [v0.15.129] - 2026-08-11 - Neese’s Cheetah
- [v0.15.128] - 2026-08-11 - Neese’s Cheetah
- [v0.15.127] - 2026-08-11 - Neese’s Cheetah
- [v0.15.126] - 2026-08-11 - Neese’s Cheetah
- [v0.15.125] - 2026-08-11 - Neese’s Cheetah
- [v0.15.124] - 2026-08-10 - Neese’s Cheetah
- [v0.15.123] - 2026-08-10 - Neese’s Cheetah
- [v0.15.122] - 2026-08-09 - Neese’s Cheetah
- [v0.15.121] - 2026-08-08 - Neese’s Cheetah
- [v0.15.120] - 2026-08-08 - Neese’s Cheetah
- [v0.15.119] - 2026-08-08 - Neese’s Cheetah
- [v0.15.118] - 2026-08-08 - Neese’s Cheetah
- [v0.15.117] - 2026-08-07 - Neese’s Cheetah
- [v0.15.116] - 2026-08-06 - Neese’s Cheetah
- [v0.15.115] - 2026-08-06 - Neese’s Cheetah
- [v0.15.114] - 2026-08-05 - Neese’s Cheetah
- [v0.15.113] — 2026-08-05 — Neese’s Cheetah
- [v0.15.112] — 2026-08-03 — Neese’s Cheetah
- [v0.15.111] - 2026-08-03 - Neese’s Cheetah
- [v0.15.110] - 2026-08-02 - Neese’s Cheetah
- [v0.15.109] - 2026-08-02 - Neese’s Cheetah
- [v0.15.108] - 2026-08-01 - Neese’s Cheetah
- [v0.15.107] - 2026-08-01 - Neese’s Cheetah
- [v0.15.106] - 2026-08-01 - Neese’s Cheetah
- [v0.15.105] - 2026-08-01 - Neese’s Cheetah
- [v0.15.104] - 2026-08-01 - Neese’s Cheetah
- [v0.15.103] - 2026-08-01 - Neese’s Cheetah
- [v0.15.102] - 2026-07-31 - Neese’s Cheetah
- [v0.15.101] - 2026-07-31 - Neese’s Cheetah
- [v0.15.100] - 2026-07-30 - Neese’s Cheetah
- [v0.15.99] - 2026-07-30 - Neese’s Cheetah
- [v0.15.98] - 2026-07-30 - Neese’s Cheetah
- [v0.15.97] - 2026-07-30 - Neese’s Cheetah
- [v0.15.96] - 2026-07-30 - Neese’s Cheetah
- [v0.15.95] - 2026-07-30 - Neese’s Cheetah
- [v0.15.94] - 2026-07-30 - Neese’s Cheetah
- [v0.15.93] - 2026-07-30 - Neese’s Cheetah
- [v0.15.92] - 2026-07-30 - Neese’s Cheetah
- [v0.15.91] - 2026-07-30 - Neese’s Cheetah
- [v0.15.90] - 2026-07-29 - Neese’s Cheetah
- [v0.15.89] - 2026-07-29 - Neese’s Cheetah
- [v0.15.88] - 2026-07-29 - Neese’s Cheetah
- [v0.15.87] - 2026-07-29 - Neese’s Cheetah
- [v0.15.86] - 2026-07-29 - Neese’s Cheetah
- [v0.15.85] - 2026-07-29 - Neese’s Cheetah
- [v0.15.84] - 2026-07-29 - Neese’s Cheetah
- [v0.15.83] - 2026-07-28 - Neese’s Cheetah
- [v0.15.82] - 2026-07-28 - Neese’s Cheetah
- [v0.15.81] - 2026-07-28 - Neese’s Cheetah
- [v0.15.80] - 2026-07-28 - Neese’s Cheetah
- [v0.15.79] - 2026-07-28 - Neese’s Cheetah
- [v0.15.78] - 2026-07-28 - Neese’s Cheetah
- [v0.15.77] - 2026-07-28 - Neese’s Cheetah
- [v0.15.76] - 2026-07-28 - Neese’s Cheetah
- [v0.15.75] - 2026-07-28 - Neese’s Cheetah
- [v0.15.74] - 2026-07-28 - Neese’s Cheetah
- [v0.15.73] - 2026-07-28 - Neese’s Cheetah
- [v0.15.72] - 2026-07-28 - Neese’s Cheetah
- [v0.15.71] - 2026-07-27 - Neese’s Cheetah
- [v0.15.70] - 2026-07-27 - Neese’s Cheetah
- [v0.15.69] - 2026-07-27 - Neese’s Cheetah
- [v0.15.68] - 2026-07-27 - Neese’s Cheetah
- [v0.15.67] - 2026-07-27 - Neese’s Cheetah
- [v0.15.66] - 2026-07-26 - Neese’s Cheetah
- [v0.15.65] - 2026-07-26 - Neese’s Cheetah
- [v0.15.64] - 2026-07-26 - Neese’s Cheetah
- [v0.15.63] - 2026-07-26 - Neese’s Cheetah
- [v0.15.62] - 2026-07-26 - Neese’s Cheetah
- [v0.15.61] - 2026-07-26 - Neese’s Cheetah
- [v0.15.60] - 2026-07-25 - Neese’s Cheetah
- [v0.15.59] - 2026-07-25 - Neese’s Cheetah
- [v0.15.58] - 2026-07-24 - Neese’s Cheetah
- [v0.15.57] - 2026-07-24 - Neese’s Cheetah
- [v0.15.56] - 2026-07-23 - Neese’s Cheetah
- [v0.15.55] - 2026-07-23 - Neese’s Cheetah
- [v0.15.54] - 2026-07-23 - Neese’s Cheetah
- [v0.15.53] - 2026-07-23 - Neese’s Cheetah
- [v0.15.52] - 2026-07-23 - Neese’s Cheetah
- [v0.15.51] - 2026-07-22 - Neese’s Cheetah
- [v0.15.50] - 2026-07-20 - Neese’s Cheetah
- [v0.15.49] - 2026-07-20 - Neese’s Cheetah
- [v0.15.48] - 2026-07-20 - Neese’s Cheetah
- [v0.15.47] - 2026-07-19
- [v0.15.46] - 2026-07-19
- [v0.15.45] - 2026-07-18 - Neese’s Cheetah
- [v0.15.44] - 2026-07-17 - Neese’s Cheetah
- [v0.15.43] - 2026-07-17 - Neese’s Cheetah
- [v0.15.42] - 2026-07-17 - Neese’s Cheetah
- [v0.15.41] - 2026-07-16 - Neese’s Cheetah
- [v0.15.40] - 2026-07-15 - Neese’s Cheetah
- [v0.15.39] - 2026-07-14 - Neese’s Cheetah
- [v0.15.38] - 2026-07-13 - Neese’s Cheetah
- [v0.15.37] - 2026-07-12 - Neese’s Cheetah
- [v0.15.36] - 2026-07-12 - Neese’s Cheetah
- [v0.15.35] - 2026-07-11 - Neese’s Cheetah
- [v0.15.34] - 2026-07-11 - Neese’s Cheetah
- [v0.15.33] - 2026-07-10 - Neese’s Cheetah
- [v0.15.32] - 2026-07-10 - Neese’s Cheetah
- [v0.15.31] - 2026-07-10 - Neese’s Cheetah
- [v0.15.30] - 2026-07-07 - Neese’s Cheetah
- [v0.15.29] - 2026-07-07 - Neese’s Cheetah
- [v0.15.28] - 2026-07-05 - Neese’s Cheetah
- [v0.15.27] - 2026-07-05 - Neese’s Cheetah
- [v0.15.26] - 2026-07-05 - Neese’s Cheetah
- [v0.15.25] - 2026-07-04 - Neese’s Cheetah
- [v0.15.24] - 2026-07-04 - Neese’s Cheetah
- [v0.15.23] - 2026-07-04 - Neese’s Cheetah
- [v0.15.22] - 2026-07-04 - Neese’s Cheetah
- [v0.15.21] - 2026-07-03 - Neese’s Cheetah
- [v0.15.20] - 2026-07-03 - Neese’s Cheetah
- [v0.15.19] - 2026-07-03 - Neese’s Cheetah
- [v0.15.18] - 2026-07-02 - Neese’s Cheetah
- [v0.15.17] - 2026-07-02 - Neese’s Cheetah
- [v0.15.16] - 2026-07-02 - Neese’s Cheetah
- [v0.15.15] - 2026-07-02 - Neese’s Cheetah
- [v0.15.14] - 2026-07-02 - Neese’s Cheetah
- [v0.15.13] - 2026-07-01 - Neese’s Cheetah
- [v0.15.12] - 2026-07-01 - Neese’s Cheetah
- [v0.15.11] - 2026-07-01 - Neese’s Cheetah
- [v0.15.10] - 2026-07-01 - Neese’s Cheetah
- [v0.15.9] - 2026-07-01 - Neese’s Cheetah
- [v0.15.8] - 2026-06-30 - Neese’s Cheetah
- [v0.15.7] - 2026-06-30 - Neese’s Cheetah
- [v0.15.6] - 2026-06-29 - Neese’s Cheetah
- [v0.15.5] - 2026-06-29 - Neese’s Cheetah
- [v0.15.4] - 2026-06-29 - Neese’s Cheetah
- [v0.15.3] - 2026-06-29 - Neese’s Cheetah
- [v0.15.2] — 2026-06-28 — Neese’s Cheetah
- [v0.15.1] — 2026-06-26 — Neese’s Cheetah
- [v0.15.0] — 2026-06-26 — Neese’s Cheetah
- [v0.14.1] — 2026-06-24 — Bartlett’s Goose
- [v0.14.0] — 2026-06-22 — Bartlett’s Goose
- [v0.13.1] — 2026-06-17
- [v0.13.0] — 2026-06-17 — Wisesa’s Fox
- [v0.12.2] — 2026-06-12
- [v0.12.1] — 2026-06-12
- [v0.12.0] — 2026-06-12 — Knuth’s Beaver
- [v0.11.4] — 2026-06-05
- [v0.11.3] — 2026-06-04
- [v0.11.2] — 2026-06-04
- [v0.11.1] — 2026-06-03
- [v0.11.0] — 2026-06-03 — Sun’s Stingray
- [v0.10.5] — 2026-06-01
- [v0.10.4] — 2026-06-01
- [v0.10.3] — 2026-05-30
- [v0.10.2] — 2026-05-29
- [v0.10.1] — 2026-05-29
- [v0.10.0] — 2026-05-27 — Pisani’s Penguin
- [v0.9.1] — 2026-05-22
- [v0.9.0] — 2026-05-22 — Knowles’s Kingfisher
- Historical Codename Artwork (pre-v0.9.0)
- Release process
- 0.18.0 candidate scope: Berkelbach’s Badger
- GitHub source publication
- Website sources and shared appearance
- Contributing to vibe-qc
- Contributor setup, fresh-clone bootstrap and rebuild discipline
- Developer Test Lanes
- Matched periodic performance reports
- What validation actually runs
- Support vibe-qc
- Sponsors
- How to cite vibe-qc
- License + bundled-data inventory
- Example scripts and generated outputs
Status
vibe-qc is pre-release software heading toward a 1.0 feature-complete milestone. Molecular HF/DFT and the documented analysis routes are stable. MP2 and canonical CCSD(T) have small-molecule numerical validation, but their production-size memory envelope remains gated by BUG 63 until bounded-memory executions, rather than dry-run estimates, are archived. The periodic stack supports three production routes: GDF (µHa parity vs PySCF), GPW (Gaussian Plane Waves with Γ-point analytic gradients; MPI grid overlay experimental), and BIPOLE (exact Ewald-J with production finite-difference forces and fixed-cell atomic relaxation; analytic gradients remain a gated preview). The eigensolver framework offers Davidson and LOBPCG by keyword, with experimental Jacobi-Davidson and GPLHR at library level. Analysis tools (COOP/COHP, QTAIM, Mayer bond orders, fat bands) and the basis_toolkit import/export system shipped in v0.15.0. The ab-initio CCM (Γ-CCM and χ-CCM) ships as experimental. Follow the roadmap for what’s next.
Licensed under the Mozilla Public License 2.0. Source at github.com/vibe-qc/vibe-qc.
Feedback and bug reports
Found a bug, have a feature request, or want to send a patch? The decision tree lives in CONTRIBUTING.md. The short version:
Bugs / install problems / feature requests → GitHub issues
Security vulnerabilities → email
mpei@vibe-qc.com(see SECURITY.md)General feedback / questions → email
mpei@vibe-qc.comSponsor the project → see Support vibe-qc (GitHub Sponsors, Ko-fi)